
In my work on solid electrolyte cell architectures, I have concentrated on the central bottleneck that prevents NASICON-type electrolytes from being deployed in practical lithium metal batteries: the thermodynamic incompatibility between the electrolyte and the lithium anode. When a lithium metal anode is placed in direct contact with a NASICON solid electrolyte such as Li1.3Al0.3Ti1.7(PO4)3 (LATP), a spontaneous reduction of Ti4+ to Ti3+ occurs, accompanied by the continuous formation of a mixed-conducting interphase. Unlike the solid electrolyte interphase in liquid systems, this interphase is not self-limiting because it conducts both ions and electrons. The result is a persistent interfacial decomposition, rising resistance, mechanical degradation, and eventually the loss of the solid electrolyte cell function. I therefore set out to design an artificial interface that simultaneously protects the electrolyte, regulates lithium deposition, and acts as an active three-dimensional lithium host rather than a passive buffer layer.
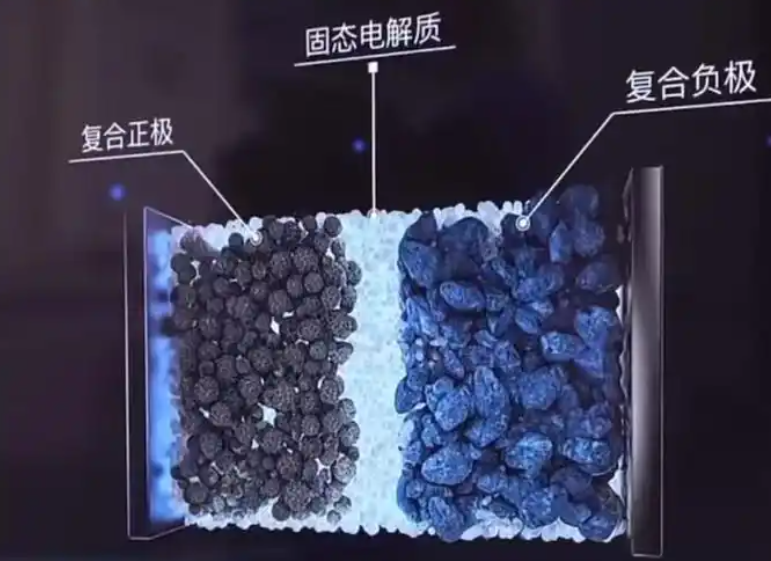
The strategy I adopted is a bilayer electron/ion mixed-conducting interface fabricated by sequential electrospinning and magnetron sputtering. From the lithium anode side toward the solid electrolyte, the bilayer comprises a lithiophilic mixed-conducting layer (MCL) and an electronically insulating but ion-conducting layer (ICL). The MCL contains single-walled carbon nanotubes and gold nanoparticles, giving it both electronic and ionic conductivity, while the ICL contains only ionic pathways. This architecture ensures that electrons cannot percolate into the NASICON electrolyte, yet lithium ions can migrate freely. Upon electrochemical activation, the MCL transforms into an in-situ three-dimensional lithium anode, and the porous fibrous scaffold accommodates lithium plating and stripping without large volume fluctuations. In this article I describe the design principles, the fabrication route, the structural and electrochemical characterization, the finite element analysis, the thermal stability evaluation, and the full-cell performance of this solid electrolyte cell concept.
Design Rationale and Comparison with Previous Interfacial Strategies
Most reported interfacial engineering approaches for NASICON electrolytes rely on two-dimensional planar coatings. These include sputtered oxide films, fluoride-based coatings, polymer buffer layers, and composite interlayers. Although such planar designs improve stability to some degree, they offer limited control over out-of-plane lithium growth and poor accommodation of volume changes during cycling. I decided to move from a two-dimensional architecture to a three-dimensional fibrous architecture because a three-dimensional host can distribute the local current density over a much larger effective surface area. Table 1 summarizes the conceptual differences between the conventional planar approach and my bilayer three-dimensional approach.
| Feature | Conventional planar interlayer | My bilayer 3D interlayer |
|---|---|---|
| Dimensionality | 2D continuous film | 3D electrospun fiber network |
| Lithium host function | Passive buffer | Active in-situ 3D anode |
| Electronic control | Usually insulating | Graded: mixed-conducting MCL + insulating ICL |
| Nucleation sites | Limited | Abundant Au nanoparticle seeds |
| Stress accommodation | Poor | High porosity buffer |
| Li+ transference | Moderate | Enhanced by solid–ionic liquid composite |
| Thermal protection | Rarely validated | Suppresses molten Li attack up to 300 °C |
The design principles I followed can be summarized as follows. First, the layer facing lithium must be lithiophilic and mixed-conducting so that it wets lithium, provides nucleation sites, and conducts both electrons and ions. Second, the layer facing the NASICON electrolyte must be electronically insulating to block electron percolation, which is the root cause of continuous reduction. Third, the entire interlayer must be porous enough to host lithium and to buffer volume changes. Fourth, the interlayer must be compatible with a small amount of ionic liquid that improves interfacial contact and ionic transport. These principles guided every step of the fabrication and characterization described below.
Fabrication of the Bilayer Interface
I prepared two separate electrospinning dispersions. The dispersion for the ICL contained polyacrylonitrile (PAN), poly(vinylidene fluoride) (PVDF), nano-LATP, and lithium perchlorate in N,N-dimethylformamide. The mass ratio of PAN, PVDF, and nano-LATP was 9:1:5, and the lithium salt was added to achieve a molar ratio of lithium ions to nitrile groups of 1:10. The total solid content was 12.5 wt%. The dispersion for the MCL was prepared in the same way except that single-walled carbon nanotubes were added together with nano-LATP before the polymers. The mass ratio of PAN, PVDF, nano-LATP, and carbon nanotubes was 9:1:5:1, and the total solid content was 10.5 wt%. Table 2 lists the two dispersion formulations.
| Component | ICL dispersion | MCL dispersion |
|---|---|---|
| PAN (g) | 9 | 9 |
| PVDF (g) | 1 | 1 |
| Nano-LATP (g) | 5 | 5 |
| Single-walled CNTs (g) | 0 | 1 |
| LiClO4 | [Li+]:[CN] = 1:10 | [Li+]:[CN] = 1:10 |
| Solvent | DMF | DMF |
| Total solid content (wt%) | 12.5 | 10.5 |
Electrospinning was performed with a needle of 0.27 mm inner diameter, an applied voltage of 30 kV, and a tip-to-drum distance of 10 cm. The drum collector rotated at 350 r min−1, and the feeding rate was 0.8 mL h−1. The ambient temperature was maintained at 30 ± 2 °C with a relative humidity of 40 ± 5%. I electrospun the ICL dispersion first for 2 h and then the MCL dispersion for 2 h, producing a bilayer with a total thickness of approximately 30 µm and roughly equal thickness contributions from the two layers. Gold nanoparticles were then deposited onto the MCL surface by magnetron sputtering under an argon atmosphere at 0.5 Pa with a current of 30 mA for 10 s. The resulting interlayer was cut into discs of 12 mm diameter and vacuum-dried at 50 °C for 72 h before use.
The NASICON solid electrolyte itself was prepared by a solid-state reaction. I mixed lithium carbonate, aluminum oxide, titanium dioxide, and ammonium dihydrogen phosphate in acetone by wet ball milling, calcined the dried precursor at 900 °C for 10 h, and then milled the resulting ceramic blocks. A coarse milling step produced powder with an average particle size of about 600 nm, and a subsequent fine milling step produced nano-LATP with an average particle size of about 100 nm. The coarse powder was uniaxially pressed into 12 mm pellets at 150 MPa and sintered at 900 °C for 6 h in air. The sintered pellets were polished and cleaned before cell assembly. This solid electrolyte cell platform allowed me to compare the bilayer interface against conventional liquid electrolyte and ionic liquid wetting in a controlled manner.
Structural and Chemical Characterization
Scanning electron microscopy revealed that the MCL consists of randomly oriented fibers with abundant inter-fiber space. Gold nanoparticles and nano-LATP fillers are uniformly distributed on the fiber surfaces. The ICL exhibits a similar fibrous structure but without gold nanoparticles, and the nano-LATP fillers are embedded within the fibers. Cross-sectional imaging confirmed the bilayer architecture with a total thickness near 30 µm. Energy-dispersive X-ray spectroscopy confirmed the homogeneous distribution of aluminum, titanium, and phosphorus from the LATP fillers as well as the presence of gold on the MCL fibers. The four-point probe measurement gave an electronic conductivity of 7.0 × 10⁻³ S cm−1 for the MCL, whereas no electronic conductivity was detected for the ICL. The porosity of the bilayer interface was determined to be as high as 89%.
The porosity can be estimated from the apparent density and the skeletal density according to
$$P = \left(1 – \frac{\rho_{\text{app}}}{\rho_{\text{skeletal}}}\right) \times 100\%$$
where P is the porosity, ρapp is the apparent density of the fibrous mat, and ρskeletal is the density of the solid constituents. A porosity of 89% means that the fibrous network provides a large internal volume for ionic liquid uptake and for lithium storage. The ionic liquid uptake was approximately 825 wt%, which matches the pore volume and ensures that the interlayer is fully wetted during cell operation.
X-ray diffraction patterns of the interfacial layers and the raw materials confirmed the crystalline structure. PAN shows characteristic peaks near 17.0° and 29.3°, corresponding to the (100) and (110) planes. PVDF shows peaks near 18.4°, 20.0°, and 26.6°, indexed to the (020), (110), and (021) planes. Carbon nanotubes display peaks near 26.0° and 43.0°, corresponding to the (002) and (100) planes of graphite. Nano-LATP shows sharp peaks matching the NASICON structure. In both the ICL and the MCL, the LATP peaks remain prominent while the polymer peaks are substantially weakened, indicating that the nano-LATP fillers reduce polymer crystallinity. This structural change is favorable for lithium ion migration because it increases the amorphous fraction and creates continuous ionic pathways along the filler surfaces. Table 3 summarizes the key structural and transport parameters of the two layers.
| Parameter | ICL | MCL |
|---|---|---|
| Electronic conductivity (S cm−1) | Not detected | 7.0 × 10⁻³ |
| Ionic conductivity (S cm−1) | ~10−4 to 10−3 | ~10−3 |
| Porosity (%) | ~89 | ~89 |
| Thickness (µm) | ~15 | ~15 |
| Gold nanoparticles | Absent | Present |
| Carbon nanotubes | Absent | Present |
| Primary function | Electron blocking and ionic conduction | Lithiophilic nucleation and mixed conduction |
Electrochemical Performance in Symmetric Solid Electrolyte Cells
I assembled symmetric solid electrolyte cells with the configuration lithium | MCL | ICL | LATP | ICL | MCL | lithium. The MCL faced the lithium metal, and a small amount of ionic liquid was added to each interlayer. For comparison, I also assembled cells wetted with a conventional liquid electrolyte and cells wetted with the same ionic liquid but without the bilayer interlayer. The symmetric cells were cycled at a current density of 0.1 mA cm−2 with an areal capacity of 0.1 mAh cm−2.
The cell with the bilayer interlayer exhibited an ultralow overpotential of approximately 10 mV and cycled for more than 1400 h. In contrast, the ionic-liquid-wetted cell failed after 842 h and the liquid-electrolyte-wetted cell failed after 683 h. The initial overpotentials of the control cells were approximately 75 mV and 80 mV, respectively, and they increased continuously during cycling. These results demonstrate that the bilayer interface accelerates lithium plating and stripping kinetics and enhances reversibility in the solid electrolyte cell. Table 4 compares the long-term cycling performance of the three configurations.
| Cell configuration | Initial overpotential (mV) | Cycling lifetime (h) | Failure mode |
|---|---|---|---|
| Bilayer interlayer | ~10 | >1400 | No failure observed |
| Ionic liquid only | ~75 | 842 | Interfacial degradation |
| Liquid electrolyte only | ~80 | 683 | Interfacial degradation and fracture |
I retrieved the LATP pellets from the different cells and examined their morphology. Pellets protected by the bilayer interface remained intact and white even after 500 and 1000 h of cycling. Pellets from the ionic-liquid-wetted and liquid-electrolyte-wetted cells fragmented severely and developed dark-purple or black reaction products on their surfaces. X-ray photoelectron spectroscopy of the Ti 2p region showed only Ti4+ peaks at 459.7 and 465.2 eV for the pristine pellet and for the bilayer-protected pellet. In contrast, additional Ti3+ peaks at 458.6 and 463.5 eV appeared for the control pellets, indicating reduction of the NASICON electrolyte. The pellet retrieved from the bilayer-protected cell after 1000 h showed no Ti3+ signal at all. This is direct evidence that the ICL prevents electron percolation and thereby suppresses the reduction of Ti4+ in the solid electrolyte cell.
Electrochemical impedance spectroscopy provided further insight. In the equivalent circuit, the resistance associated with lithium ion migration through the interphase and the resistance associated with the interface between the interlayer and the LATP pellet both increased rapidly for the control cells. For the bilayer-equipped cell, the interphase resistance remained at approximately 7 Ω cm2 per side and the interlayer–LATP interface resistance increased only gradually from about 13 to 20 Ω cm2. The formation of a highly conductive and chemically stable interphase explains the low and stable polarization.
The lithium ion transference number was determined by the Bruce–Vincent method using
$$t_{\mathrm{Li}^+} = \frac{I_s(\Delta V – I_0 R_0)}{I_0(\Delta V – I_s R_s)}$$
where I0 and Is are the initial and steady-state currents, ΔV is the applied potential, and R0 and Rs are the charge transfer resistances before and after polarization. The bilayer interface gave a transference number of 0.60, compared with 0.21 for the ionic-liquid-wetted interface and 0.20 for the liquid-electrolyte-wetted interface. The high transference number originates from the porous solid–ionic liquid composite architecture. The confinement of the ionic liquid within the fibrous network restricts long-range anion migration, the embedded nano-LATP fillers provide solid-assisted lithium ion pathways, the mixed-conducting MCL directs a preferential lithium ion flux toward the lithiophilic gold nanoparticles, and the interconnected structure shortens local diffusion paths. Table 5 summarizes the transference numbers and the interfacial resistances.
| Interfacial system | Li+ transference number | Interphase resistance (Ω cm2) | Interlayer–LATP resistance (Ω cm2) |
|---|---|---|---|
| Bilayer interlayer | 0.60 | ~7 | 13–20 |
| Ionic liquid only | 0.21 | Rapidly increasing | Rapidly increasing |
| Liquid electrolyte only | 0.20 | Rapidly increasing | Rapidly increasing |
Interfacial Evolution and Lithium Deposition Regulation
The cycling-induced interfacial evolution differs dramatically between the bilayer-protected solid electrolyte cell and the control cells. In the liquid-electrolyte-wetted cell, internal pressure partially expels the liquid from the interface, and severe side reactions occur between LATP and lithium. The pellet fractures, discolors, and develops a rough lithium surface with protrusions. In the bilayer-protected cell, the fibrous interlayer provides ample free space for capillary-driven ionic liquid infiltration and stable retention. During activation, the MCL decorated with high-surface-area gold nanoparticles provides abundant nucleation sites and lithium accommodation space, while its mixed conductivity guides uniform lithium deposition. The ICL physically isolates lithium from LATP and confers chemical and electrochemical stability. As activation proceeds, the interface evolves into a robust three-dimensional mixed-conducting lithium anode, enabling cooperative lithium plating and stripping across the MCL and the lithium surface.
I examined the lithium anodes retrieved from the different cells. After 500 h of cycling, the bilayer-protected lithium anode retained a smooth and intact metallic surface, nearly identical to pristine lithium. Scanning electron microscopy revealed a dense, moss-like morphology characterized by uniform lithium deposition along the MCL fibers. Energy-dispersive X-ray spectroscopy confirmed a homogeneous distribution of fluorine, indicating uniform interphase coverage. Carbon and oxygen signals originated mainly from surface byproducts formed during brief air exposure. The ICL side exhibited no lithium deposition and only a slight thickness increase due to swelling. Even after 1000 h of cycling, the three-dimensional anode retained a flat surface and interphase integrity. In contrast, the liquid-electrolyte-wetted lithium anode displayed severe roughening, protrusion growth, and substantial dead lithium accumulation after 500 h, accompanied by uneven interphase reconstruction and LATP-derived fragments identified by the titanium signal. Dendritic growth was also observed in the ionic-liquid-wetted cells, confirming that unregulated lithium ion flux inevitably triggers local plating.
X-ray photoelectron spectroscopy with depth profiling revealed the chemical composition of the interphase on the lithium anode as a function of etching time. The breakdown of the ionic liquid and the lithium salt generated abundant lithium fluoride at 685 eV throughout the interphase layer on the MCL side. Lithium fluoride is known to facilitate the formation of a highly ion-conducting interphase. Comparison of the results after 500 and 1000 h of cycling showed that the elemental composition remained relatively stable, reflecting long-term compositional stability. In the control cells, lithium fluoride was also detected but its content was limited and its distribution was uneven, so it failed to provide effective interfacial protection. Table 6 summarizes the interphase composition on the three-dimensional anode.
| Species | Binding energy (eV) | Role in the interphase |
|---|---|---|
| LiF | 685 | Ion-conducting, mechanically robust |
| Li2CO3 | ~289 | Surface byproduct from air exposure |
| LixPOyFz | ~134–136 | Phosphorus-containing interphase component |
| Li–C–O | ~531 | Organic and inorganic carbonates |
| Ti3+ | 458.6 and 463.5 | Degradation indicator for LATP |
Finite Element Simulation of Lithium Deposition
To visualize the electric field distribution, the lithium ion concentration field, and the thickness evolution of the lithium anode during deposition, I established an electrodeposition model using finite element analysis. The model solves the tertiary current distribution, electrode surface deformation, electric field distribution, ion migration, and concentration variations. The electrodeposition kinetics are described by the Butler–Volmer equation
$$j = j_0 \left[ \exp\left(\frac{\alpha_a F \eta}{RT}\right) – \exp\left(-\frac{\alpha_c F \eta}{RT}\right) \right]$$
where j is the current density, j0 is the exchange current density, αa and αc are the anodic and cathodic charge transfer coefficients, F is the Faraday constant, R is the universal gas constant, T is the absolute temperature, and η is the overpotential. Under lithium deposition conditions, the electrode is negatively polarized and the cathodic reduction reaction dominates, so the anodic term can be neglected and the equation simplifies to
$$j = j_0 \exp\left(-\frac{\alpha_c F \eta}{RT}\right)$$
Because the reduction of lithium ions at the electrode exhibits relatively fast electrochemical kinetics, the overall deposition process is predominantly governed by lithium ion diffusion from the bulk electrolyte to the electrode surface, which follows Fick’s second law
$$\frac{\partial C_{\mathrm{Li}}(x,t)}{\partial t} = D_{\mathrm{Li}} \frac{\partial^2 C_{\mathrm{Li}}(x,t)}{\partial x^2}$$
where CLi(x,t) is the lithium ion concentration at position x above the electrode surface and time t, and DLi is the diffusion coefficient of lithium ions. The concentration overpotential arising from local lithium ion depletion near the electrode interface is expressed by the modified Nernst equation
$$\eta = \frac{RT}{F} \ln\left(\frac{C_{\mathrm{Li}}(0,t)}{C_{\mathrm{Li,bulk}}}\right)$$
where CLi,bulk is the bulk lithium ion concentration in the electrolyte and CLi(0,t) represents the concentration at the electrode surface. Ionic current conservation and concentration-dependent transport were solved simultaneously using the tertiary current distribution module to capture the coupled effects of electrochemical reaction kinetics and ion transport under non-uniform interfacial conditions.
In the two-dimensional model, the electrodes were set to 27 µm in length and 20 µm in width, with a randomly generated initial geometry on the lithium anode side. In the control group, lithium protrusion growth was modeled based on theoretical values and the protruding structures observed in scanning electron microscopy. In the experimental group, the diameters of the initial fibrous structures were determined by statistical analysis of the microscopy images. Model parameters, including the initial lithium ion concentration, the exchange current density, and the lithium ion diffusion coefficient, were obtained from experimental measurements. Lithium deposition was simulated under potentiostatic conditions with the anode surface potential fixed at 0 V and a constant voltage of 0.2 mV applied at the top boundary. Table 7 lists the key simulation parameters.
| Parameter | Value | Source |
|---|---|---|
| Initial Li+ concentration | 1.0 mol L−1 | Electrolyte formulation |
| Exchange current density | 1.0 × 10⁻³ A cm−2 | Tafel analysis |
| Li+ diffusion coefficient (bilayer) | 1.23 × 10⁻⁹ cm2 s−1 | Warburg analysis |
| Li+ diffusion coefficient (liquid electrolyte) | 7.74 × 10⁻¹¹ cm2 s−1 | Warburg analysis |
| Li+ diffusion coefficient (ionic liquid) | 8.52 × 10⁻¹¹ cm2 s−1 | Warburg analysis |
| Anode surface potential | 0 V | Boundary condition |
| Top boundary voltage | 0.2 mV | Boundary condition |
The simulation revealed that at the liquid-electrolyte-wetted interface, surface roughness of the lithium anode induces a non-uniform local electric field, which drives lithium ion accumulation at microscopic irregular tips and forms steep electric field gradients. Sluggish lithium ion transport in the liquid electrolyte leads to gradual ion depletion near the anode and the development of significant concentration gradients, which in turn elevate the local current density and trigger non-uniform lithium deposition. The combined effects of the electric field and concentration gradients ultimately trigger the formation of lithium protrusions. These simulation results align with the experimental observations of significant surface roughening and dendritic features in the control cells.
In contrast, the interlayer-built three-dimensional anode benefits from the synergistic facilitation of lithium ion transport by the solid–ionic liquid composite interlayer, which mitigates concentration gradients. Lithium deposition is observed exclusively on the MCL, and the in-situ three-dimensional anode develops a uniform moss-like morphology. Analysis of surface thickness evolution during plating provides further insight into interphase integrity. At the liquid-electrolyte-wetted interface, the lithium anode undergoes pronounced volumetric expansion, with certain surface regions stretched and prone to interphase rupture, while other regions thicken markedly due to lithium metal deposition. In the three-dimensional anode, lithium deposits cooperatively across the large effective surface area of the MCL fibers and the lithium anode, producing evenly distributed thickness changes with only slight surface thickening. This minimizes local volume variations and preserves interphase integrity, which is corroborated by the experimental observation of a continuous and compositionally stable interphase on the in-situ three-dimensional anode.
Comparative Experiments on Functional and Thermal Stability
To evaluate the role of the artificial interlayer in preserving the structural integrity of the NASICON pellet, I conducted a puncture experiment. In symmetric solid electrolyte cells with the same configuration described earlier, I introduced a micropuncture on one side of the interlayer while leaving the remaining regions intact. The cells were cycled at 0.1 mA cm−2 and 0.1 mAh cm−2. Pronounced asymmetric overpotential fluctuations appeared within 200 h of cycling. The optical image of the pellet showed a purple-degraded region solely on the punctured side, corresponding to the damaged area. Scanning electron microscopy revealed pronounced surface roughening in the degraded region, whereas the interlayer-protected regions remained white and structurally intact. Energy-dispersive X-ray spectroscopy showed substantially higher carbon content in the degraded region, resulting from air exposure of lithium, which indirectly confirms lithium dendrite penetration in the unprotected region.
To evaluate the thermal stability of the interlayer, I heated samples at 300 °C in an argon-filled box, which is above the onset temperature of the violent self-heating reaction between LATP and lithium reported in previous studies. I compared an LATP pellet in direct contact with lithium against an LATP pellet protected by the interlayer placed between the pellet and the lithium. The unprotected pellet began to crack at 31 s, suffered severe fragmentation at 36 s, and underwent violent reaction with thermal runaway accompanied by spark emission at 37 s. By 39 s, the fragments remained red-hot but the flames gradually extinguished, and the pellet ultimately turned dark purple. This thermal runaway of the NASICON electrolyte under extreme heating poses fatal risks for any practical solid electrolyte cell application and must be strictly avoided.
In contrast, the LATP pellet protected by the interlayer showed no violent reaction throughout the heating process, producing only substantial black reaction products around it. These black products likely originate from amorphous carbon formed by the decomposition of the ionic liquid at elevated temperatures. This carbon layer effectively insulates the LATP pellet from molten lithium and thereby mitigates potential thermal runaway. Importantly, the interlayer-protected pellet retained its structural integrity and functionality after heating, without any short circuiting. The retrieved pellet was cleaned and reassembled into a lithium symmetric solid electrolyte cell with new interlayers, and the cell still exhibited stable cycling at 0.1 mA cm−2. Table 8 compares the thermal behavior of the protected and unprotected configurations.
| Observation | LATP in direct contact with Li | LATP protected by bilayer |
|---|---|---|
| Onset of visible cracking | 31 s | Not observed |
| Severe fragmentation | 36 s | Not observed |
| Spark emission / thermal runaway | 37 s | Not observed |
| Final appearance | Dark purple fragments | Intact pellet with black carbon residue |
| Post-heating cell function | Destroyed | Stable cycling retained |
To clarify the functional role of each interfacial component, I also evaluated symmetric solid electrolyte cells containing LATP pellets with a bilayer interface but without gold nanoparticles, with only an MCL single-layer interface, and with only an ICL single-layer interface. The results showed that the bilayer interface could not be efficiently activated in the absence of gold nanoparticles, and unstable lithium plating and stripping behavior accompanied by noticeable voltage fluctuations was observed. For the cell with only an MCL single-layer interface, an unstable activation process occurred and the overpotential increased rapidly due to the lack of ICL protection, indicating severe degradation of the NASICON electrolyte. In contrast, the cell with only an ICL single-layer interface failed to achieve effective activation and exhibited a relatively high initial overpotential. Moreover, due to the absence of MCL regulation of lithium deposition, sudden drops in overpotential were frequently observed during cycling, suggesting the growth of lithium protrusions within the ICL and the occurrence of a soft short circuit. Table 9 summarizes the distinct and complementary roles of the components.
| Component | Primary function | Consequence if absent |
|---|---|---|
| Gold nanoparticles | Lithiophilic nucleation seeds, reduced nucleation barrier | Difficult activation, unstable plating |
| Mixed-conducting layer (MCL) | Electron and ion transport, 3D lithium host | No regulation of deposition, protrusion growth |
| Ion-conducting layer (ICL) | Electron blocking, chemical protection of LATP | Continuous reduction of Ti4+, electrolyte degradation |
| Ionic liquid | Interfacial wetting, ionic transport | Poor contact, high resistance |
| Carbon nanotubes | Electronic percolation in the MCL | Insufficient mixed conduction |
| Nano-LATP fillers | Solid-assisted Li+ pathways, reduced crystallinity | Lower ionic conductivity |
Full-Cell Performance and Electrochemical Stability Window
I evaluated the interlayer in full solid electrolyte cells composed of a lithium metal anode, a LATP pellet, and a LiFePO4 cathode. The interlayer was placed between the lithium anode and the LATP pellet, and the cathode was wetted with an ionic liquid. Before full-cell testing, I measured the electrochemical stability window by linear sweep voltammetry in a lithium | interlayer | stainless steel asymmetric cell at a scan rate of 0.1 mV s−1 from −0.1 to 6.0 V. The cell equipped with the interlayer exhibited well-defined lithium plating and stripping redox peaks near 0 V, while anodic current was only observed above 5.5 V, with no additional side-reaction peaks within this voltage range. This indicates that the interlayer maintains a wide and stable electrochemical stability window from 0 to 5.5 V, which fully encompasses the working potential range of the full cell and demonstrates its potential for pairing lithium anodes with high-voltage cathodes.
For the critical current density evaluation, symmetric solid electrolyte cells were subjected to galvanostatic cycling with stepwise increasing current densities from 0.1 to 2.0 mA cm−2. Cells with the ionic-liquid-wetted interface showed a voltage increase followed by an abrupt drop at 0.5 mA cm−2, while cells with the liquid-electrolyte-wetted interface showed a similar drop at 0.4 mA cm−2. These current densities were defined as the critical current densities for the respective configurations. In contrast, cells equipped with the bilayer interlayer displayed stable plating and stripping even at high current densities up to 2.0 mA cm−2. Before activation, the interlayer-equipped cell exhibited a critical current density of only 1.0 mA cm−2 together with a relatively high polarization voltage, indicating that the activation process is essential for fully establishing favorable interfacial kinetics. Long-term cycling of the interlayer-equipped symmetric solid electrolyte cell at a high current density of 1.0 mA cm−2 demonstrated stable operation for over 650 h. This operating current density approaches the critical benchmark for high-energy-density lithium metal batteries, highlighting its practical significance.
Analysis of lithium ion transport across the various interfacial systems revealed that the interlayer exhibits a diffusion coefficient of 1.23 × 10⁻⁹ cm2 s−1, approximately one and a half orders of magnitude higher than those of the liquid-electrolyte-wetted and ionic-liquid-wetted interfaces, which were 7.74 × 10⁻¹¹ and 8.52 × 10⁻¹¹ cm2 s−1, respectively. The diffusion coefficient was calculated from the low-frequency region of the impedance spectra using the Warburg impedance
$$Z_W = \sigma_W \omega^{-1/2}$$
where σW is the Warburg coefficient and ω is the angular frequency. According to Fick’s law of diffusion, the lithium ion diffusion coefficient in symmetric cells can be calculated using
$$D_{\mathrm{Li}} = \frac{R^2 T^2}{2 A^2 n^4 F^4 C^2 \sigma_W^2}$$
where R is the gas constant, T is the absolute temperature, A is the electrode area, n is the number of transferred electrons, F is the Faraday constant, C is the lithium ion concentration, and σW is the Warburg coefficient. This relationship provides a practical approach for estimating lithium ion diffusion behavior based on impedance measurements. The much higher diffusion coefficient of the bilayer system confirms that the solid–ionic liquid composite interface accelerates lithium ion transport in the solid electrolyte cell.
In full cells, the interlayer exhibited excellent electrochemical performance. Rate capability tests showed a high specific capacity of 166 mAh g−1 at 0.1 C and a capacity of 126 mAh g−1 even at 1.5 C, alongside low polarization. Long-term cycling at 0.2 C demonstrated excellent cyclability: the interlayer-equipped full cell delivered an initial capacity of 162 mAh g−1 and retained 95.5% of its capacity after 100 cycles with a Coulombic efficiency approaching 99.9%. In contrast, cells with the liquid-electrolyte-wetted interface experienced drastic capacity loss, reaching 60.0% after 100 cycles. The evolution of voltage hysteresis further confirmed the interlayer stability: the hysteresis increased only gradually with minimal fluctuations, attributed to a stable lithium metal interface and low accumulation of inactive side products. In contrast, control cells exhibited a rapid increase in hysteresis and interfacial resistance, indicative of severe interface degradation. At 0.6 C, the interlayer-equipped full cell delivered an initial capacity of 150.0 mAh g−1 and retained 95.0% of its capacity after 100 cycles. The control cell with the liquid-electrolyte-wetted interface short-circuited after only 41 cycles. Table 10 summarizes the full-cell performance metrics.
| Metric | Bilayer interlayer | Liquid electrolyte only |
|---|---|---|
| Initial capacity at 0.2 C (mAh g−1) | 162 | Comparable initial value |
| Capacity retention at 0.2 C after 100 cycles | 95.5% | 60.0% |
| Coulombic efficiency | 99.9% | Lower and fluctuating |
| Initial capacity at 0.6 C (mAh g−1) | 150.0 | Not stable |
| Capacity retention at 0.6 C after 100 cycles | 95.0% | Short circuit after 41 cycles |
| Rate capability at 1.5 C (mAh g−1) | 126 | Poor |
| Voltage hysteresis evolution | Gradual and minimal | Rapid increase |
The cumulative plated capacity and maximum achievable current density of symmetric cells, together with the capacity utilization of full cells at 0.1 C, are recognized as key metrics for evaluating the efficiency and practical applicability of a solid electrolyte cell. Compared with recently reported NASICON-type solid-state batteries incorporating artificial interlayers, the bilayer interface I designed delivers superior performance across these metrics. The rationally designed bilayer interlayer enables synergistic electron and ion mixed conductivity, effectively regulates lithium deposition to suppress protrusion formation, and significantly enhances lithium ion transport via the solid–ionic liquid composite interface. The low-cost and scalable fabrication process provides practical feasibility for industrial applications. Moreover, the ultra-low content of gold nanoparticles and the minimal amount of ionic liquid employed in the interlayer contribute to improved sustainability and reduced environmental impact.
Discussion of Transport Mechanisms
The transport mechanisms in the bilayer interface can be understood by considering the different conduction paths. In the MCL, electrons flow through the carbon nanotube network and the gold nanoparticles, while lithium ions migrate through the polymer matrix, along the nano-LATP filler surfaces, and through the ionic liquid confined in the pores. In the ICL, electrons are blocked because neither the polymer nor the nano-LATP provides a percolating electronic pathway, but lithium ions migrate through the same ionic pathways. This asymmetry is the key to protecting the NASICON electrolyte from reduction while maintaining ionic contact. The gold nanoparticles lower the nucleation overpotential for lithium deposition, which is described by classical nucleation theory as
$$\Delta G^* = \frac{16 \pi \gamma^3 V_m^2}{3 (n F \eta)^2}$$
where ΔG* is the nucleation energy barrier, γ is the interfacial energy, Vm is the molar volume, n is the number of electrons transferred, F is the Faraday constant, and η is the overpotential. A lithiophilic surface reduces the interfacial energy and therefore lowers the nucleation barrier, promoting uniform nucleation rather than localized deposition. The three-dimensional fibrous architecture further distributes the local current density according to
$$i_{\text{local}} = \frac{I_{\text{total}}}{A_{\text{eff}}}$$
where ilocal is the local current density, Itotal is the total applied current, and Aeff is the effective electroactive area. Because the fibrous MCL has a high specific surface area, Aeff is much larger than the geometric area, so ilocal is much smaller than it would be on a planar electrode. This is the fundamental reason why the bilayer interface suppresses dendrite formation and enables stable cycling at high current density in the solid electrolyte cell.
The ionic liquid confined in the porous network also plays a critical role. Its confinement restricts long-range anion migration, which increases the lithium ion transference number. The interaction between the ionic liquid and the polymer matrix can be described by the Vogel–Tammann–Fulcher equation for ionic conductivity
$$\sigma(T) = \sigma_0 \exp\left(-\frac{B}{T – T_0}\right)$$
where σ(T) is the ionic conductivity, σ0 is a pre-exponential factor, B is a fragility parameter, and T0 is the Vogel temperature. The amorphous character of the polymer–salt–filler composite lowers T0 and enhances ionic conductivity, which is consistent with the reduced polymer crystallinity observed by X-ray diffraction. The combination of a high transference number, a high diffusion coefficient, and a large effective area explains the excellent rate capability and long cycle life of the solid electrolyte cell.
Safety Implications and Practical Relevance
Safety is a decisive factor for the commercialization of solid electrolyte cells. The thermal stability experiment demonstrated that the bilayer interlayer suppresses the violent reaction between LATP and molten lithium up to 300 °C. This is a significant advance because thermal runaway in solid-state batteries can be triggered by mechanical, electrical, or thermal abuse. In conventional lithium-ion batteries, thermal runaway is typically initiated by short-circuiting and exothermic decomposition of the cathode and the liquid electrolyte. In solid electrolyte cells based on NASICON electrolytes, the situation is different: the solid electrolyte itself is non-flammable, but the lithium metal anode can melt at 180.5 °C, and molten lithium accelerates the decomposition of the NASICON electrolyte upon contact, accompanied by oxygen release. The violent exothermic reaction between the released oxygen and molten lithium further exacerbates the risk. The interlayer prevents direct contact between molten lithium and the NASICON electrolyte, and the carbonaceous decomposition products of the ionic liquid act as a physical barrier. This dual protection mechanism is essential for practical solid electrolyte cell deployment.
The puncture experiment further confirmed the robustness of the interlayer. When a micropuncture was intentionally introduced, only the punctured region degraded, while the protected regions remained intact. This localized failure behavior is important because it means that a small defect does not propagate across the entire interface. In contrast, a defect in a planar interlayer would allow lithium to penetrate directly into the solid electrolyte, leading to rapid cell failure. The three-dimensional fibrous architecture provides redundant pathways for ion transport, so even if some fibers are damaged, the overall ionic conductivity is maintained.
From a manufacturing perspective, the electrospinning and magnetron sputtering processes are scalable and compatible with roll-to-roll production. The use of low-cost PAN, PVDF, and nano-LATP, combined with a minimal amount of gold and ionic liquid, keeps the material cost low. The fabrication is performed at moderate temperatures and does not require expensive vacuum equipment beyond the sputtering step, which is already widely used in the battery industry. These factors make the bilayer interface an attractive candidate for practical solid electrolyte cell manufacturing.
Comparison with Alternative Interfacial Designs
I compared my bilayer interface with several alternative designs reported in the literature for NASICON-based solid electrolyte cells. Table 11 summarizes the comparison in terms of architecture, function, and performance. The key advantage of my design is the combination of a mixed-conducting layer and an ion-conducting layer in a single three-dimensional architecture. This combination provides simultaneous control over electron blocking, ion transport, lithium nucleation, and stress accommodation. Most alternative designs address only one or two of these functions. For example, a simple oxide coating can block electrons but does not provide a lithium host or regulate nucleation. A polymer buffer can accommodate stress but may not sufficiently block electron percolation. A carbon-based interlayer can provide electronic conductivity but can also accelerate electrolyte reduction if it directly contacts the NASICON surface. My bilayer design resolves these trade-offs by spatially separating the functions.
| Design type | Electron blocking | Li host | Nucleation control | Stress buffering | Thermal protection |
|---|---|---|---|---|---|
| Planar oxide coating | Yes | No | Limited | Limited | Limited |
| Polymer buffer layer | Partial | No | No | Yes | Limited |
| Carbon-based interlayer | No | Yes | Partial | Yes | Limited |
| Fluoride coating | Yes | No | Partial | Limited | Partial |
| My bilayer 3D interface | Yes | Yes | Yes | Yes | Yes |
Another important difference is the in-situ formation of the three-dimensional anode. In my design, the MCL is not merely a protective layer; it becomes part of the anode during electrochemical activation. Lithium deposits within the porous MCL, and the MCL remains electronically connected to the current collector. This in-situ anode architecture reduces the effective current density at the lithium metal surface and minimizes the volume changes that cause interphase rupture. In contrast, planar interlayers remain as separate layers and do not participate in lithium storage. The in-situ anode concept is therefore a fundamental advance in the design of solid electrolyte cell interfaces.
Remaining Challenges and Future Directions
Despite the promising results, several challenges remain. First, the long-term compatibility of the ionic liquid with the lithium metal anode and the NASICON electrolyte needs further study. Although the interlayer suppresses degradation over 1400 h, longer cycling times may reveal slow reactions. Second, the gold nanoparticles, although used in very small amounts, contribute to cost. Alternative lithiophilic materials such as silver, zinc, or bismuth could be explored. Third, the mechanical properties of the bilayer interface under high stack pressure need to be optimized. The porosity that enables lithium storage also reduces the mechanical modulus, so a balance must be found. Fourth, the scalability of the electrospinning process for large-format cells requires further development. Fifth, the thermal stability test was performed at 300 °C, but higher temperatures or prolonged exposure may reveal additional degradation pathways.
Future work should focus on several directions. In-situ characterization techniques such as operando microscopy and synchrotron-based spectroscopy could provide deeper insight into the dynamic evolution of the interface during cycling. Machine learning and high-throughput screening could accelerate the discovery of optimal material combinations for the mixed-conducting and ion-conducting layers. Multiscale modeling that couples atomistic simulations with continuum models could predict the long-term behavior of the solid electrolyte cell. Finally, the integration of the bilayer interface with high-voltage cathodes and high-capacity anodes should be demonstrated in practical pouch cells.
Conclusion
I have designed, fabricated, and characterized a bilayer electron/ion mixed-conducting interface for NASICON-based solid electrolyte cells. The interface consists of a lithiophilic mixed-conducting layer and an electronically insulating ion-conducting layer, fabricated by sequential electrospinning and magnetron sputtering. The three-dimensional fibrous architecture provides a high specific surface area, abundant nucleation sites, and a large internal volume for lithium storage. The mixed-conducting layer transforms into an in-situ three-dimensional anode during electrochemical activation, while the ion-conducting layer prevents electron percolation into the NASICON electrolyte and suppresses the reduction of Ti4+. The interface regulates lithium deposition, homogenizes the lithium ion flux, and accommodates volume changes during cycling.
The electrochemical results demonstrate that the bilayer interface enables ultralow overpotential of approximately 10 mV at 0.1 mA cm−2 for more than 1400 h in symmetric solid electrolyte cells, stable cycling at 1.0 mA cm−2 for over 650 h, and excellent full-cell performance with 95.5% capacity retention at 0.2 C and 95.0% at 0.6 C after 100 cycles. The lithium ion transference number reaches 0.60, and the lithium ion diffusion coefficient is 1.23 × 10⁻⁹ cm2 s−1, approximately one and a half orders of magnitude higher than conventional liquid or ionic liquid interfaces. Finite element simulations confirm that the three-dimensional architecture homogenizes the electric field and the lithium ion concentration field, suppressing dendrite formation. The interlayer also provides exceptional thermal stability, suppressing the violent reaction between LATP and molten lithium up to 300 °C. This work establishes a scalable and effective strategy for interfacial engineering in NASICON-based solid electrolyte cells and provides a practical route toward high-performance, safe, and durable solid-state lithium metal batteries.
In my view, the transition from planar two-dimensional interfaces to three-dimensional mixed-conducting architectures represents a promising direction for the future development of solid electrolyte cells. The rational design of multifunctional interfaces must account for multiple physicochemical factors simultaneously: chemical stability, ionic and electronic transport, mechanical robustness, thermal safety, and manufacturability. The bilayer interface described here addresses all of these factors and demonstrates that a well-designed artificial interface can transform a reactive and unstable solid electrolyte–lithium interface into a stable and high-performance solid electrolyte cell. I believe that this strategy can be extended to other NASICON-type electrolytes, garnet electrolytes, sulfide electrolytes, and even hybrid solid–liquid systems, accelerating the practical deployment of next-generation energy storage technologies.